Linjun Wang Theoretical Chemistry Group
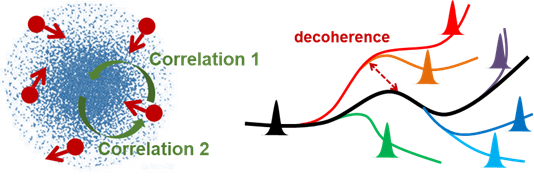 When classical nuclei interact with quantum electrons, complex quantum-classical correlation arises which cannot be simply described by the Born-Oppenheimer molecular dynamics. How to generalize adiabatic molecular dynamics into nonadiabatic cases is one of the main challenges in the quantum dynamics community. We develop mixed quantum-classical methods to realize an accurate treatment of the entangled nuclei-electron dynamics. We are also interested in exploring the underlying theoretical foundation of mixed quantum-classical trajectories that provides a systematic framework for the future development of nonadiabatic dynamics. References: 1. Jiabo Xu and Linjun Wang,* Branching Corrected Surface Hopping: Resetting Wavefunction Coefficients Based on Judgement of Wave Packet Reflection, J. Chem. Phys. 150 (16), 164101 (2019). 2. Jiabo Xu and Linjun Wang,* Branching Corrected Mean Field Method for Nonadiabatic Dynamics, J. Phys. Chem. Lett. 11 (19), 8283-8291 (2020). 3. Cancan Shao, Jiabo Xu, and Linjun Wang*, Branching and Phase Corrected Surface Hopping: A Benchmark of Nonadiabatic Dynamics in Multilevel Systems, J. Chem. Phys. 154 (23), 234109 (2021). 4. Bing Li,# Jiabo Xu,# Guijie Li, Zhecun Shi, and Linjun Wang,* A Mixed Deterministic-Stochastic Algorithm of the Branching Corrected Mean Field Method for Nonadiabatic Dynamics, J. Chem. Phys. 156 (11), 114116 (2022). 5. Guijie Li, Cancan Shao, Jiabo Xu, and Linjun Wang,* A Unified Framework of Mixed Quantum-Classical Dynamics with Trajectory Branching, J. Chem. Phys. 157 (21), 214102 (2022). 6. Guijie Li, Zhecun Shi, Xin Guo, and Linjun Wang,* What is Missing in the Mean Field Description of Spatial Distribution of Population? Important Role of Auxiliary Wave Packets in Trajectory Branching, J. Phys. Chem. Lett. 14 (44), 9855-9863 (2023). 7. Xin Guo,# Guijie Li,# Zhecun Shi, and Linjun Wang,* Surface Hopping with Reliable Wave Function by Introducing Auxiliary Wave Packets to Trajectory Branching, J. Phys. Chem. Lett. 15 (12), 3345-3353 (2024). 8. Jiabo Xu,# Zhecun Shi,# and Linjun Wang,* Consistent Construction of the Density Matrix from Surface Hopping Trajectories, J. Chem. Theory Comput. 20 (6), 2349-2361 (2024). 9. Lei Huang, Zhecun Shi, and Linjun Wang,* Detailed Complementary Consistency: Wave Function Tells Particle How to Hop, Particle Tells Wave Function How to Collapse. J. Phys. Chem. Lett. 15 (26), 6771-6781 (2024). 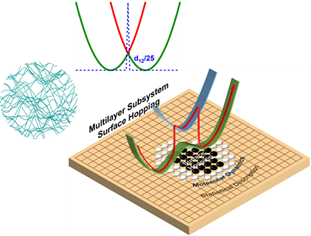 Extended condensed-phase problems often evolve complex coupling of an exponentially large number of electron and phonon degrees of freedom. Furthermore, extended systems often undergo complex energy level crossings accompanied by divergent nonadiabatic coupling, posing challenges for traditional surface hopping methods in the adiabatic representation. We aim to develop efficient surface hopping dynamics methods and employ multiscale treatments to simulate extended systems with thousands to millions of molecular sites. References: 1. Xin Bai, Jing Qiu, and Linjun Wang,* An Efficient Solution to the Decoherence Enhanced Trivial Crossing Problem in Surface Hopping, J. Chem. Phys. 148 (10), 104106 (2018). 2. Jing Qiu, Xin Bai, and Linjun Wang,* Crossing Classified and Corrected Fewest Switches Surface Hopping,J. Phys. Chem. Lett. 9 (15), 4319-4325 (2018). 3. Jing Qiu, Xin Bai, and Linjun Wang,* Subspace Surface Hopping with Size-Independent Dynamics, J. Phys. Chem. Lett. 10 (3), 637-644 (2019). 4. Jing Qiu, Yao Lu, and Linjun Wang,* Multilayer Subsystem Surface Hopping Method for Large-Scale Nonadiabatic Dynamics Simulation with Hundreds of Thousands of States, J. Chem. Theory Comput. 18 (5), 2803-2815 (2022). 5. Linjun Wang,* Jing Qiu, Xin Bai, and Jiabo Xu, Surface Hopping Methods for Nonadiabatic Dynamics in Extended Systems, WIREs Comput. Mol. Sci. 10 (2), e1435 (2020). 6. Tenghui Li, Jiawei Dong, Zihan Liu, Xiaogang Peng, and Linjun Wang,* Phonon Bottleneck in the Hot Electron Relaxation of n-Doped Quantum Dots: A Large-Scale Nonadiabatic Dynamics Perspective, ACS Nano 20 (7), 5662-5674 (2026).  Low-dimensional materials, such as one-dimensional graphene nanoribbons (GNRs) and two-dimensional transition metal dichalcogenides (TMDCs) have attracted significant attention due to their unique properties and potential applications in various fields. To effectively describe these extended systems, we employ machine learning methods to construct the condensed-phase diabatic Hamiltonian. By integrating these methods with the general non-adiabatic dynamics simulation software, SPADE (Simulation Package for non-Adiabatic Dynamics in Extended systems) developed by our group, we can simulate both theoretical models and realistic systems with high efficiency and reliability. Through our simulations and analyses, we aim to gain a deeper understanding of the electronic behavior and transport properties within these systems, with the ultimate goal of advancing the development of novel electronic and optoelectronic devices. References: 1. Xin Bai, Xin Guo, and Linjun Wang,* Machine Learning Approach to Calculate Electronic Couplings between Quasi-diabatic Molecular Orbitals: The Case of DNA, J. Phys. Chem. Lett. 12 (42), 10457-10464 (2021). 2. Zedong Wang,# Jiawei Dong,# Jing Qiu, and Linjun Wang,* All-Atom Nonadiabatic Dynamics Simulation of Hybrid Graphene Nanoribbons Based on Wannier Analysis and Machine Learning, ACS Appl. Mater. Interfaces 14 (20), 22929-22940 (2022). 3. Suryoday Prodhan,* Jing Qiu, Matteo Ricci, Otello M. Roscioni, Linjun Wang,* and David Beljonne,* Design Rules to Maximize Charge-Carrier Mobility along Conjugated Polymer Chains, J. Phys. Chem. Lett. 11 (16), 6519-6525 (2020). 4. Zirui Wang,# Jiawei Dong,# and Linjun Wang,* Large-Scale Surface Hopping Simulation of Charge Transport in Hexagonal Molecular Crystals: Role of Electronic Coupling Signs, J. Phys.: Condens. Matter 35 (34), 345401 (2023). 5. Jiawei Dong, Jing Qiu, Xin Bai, Zedong Wang, Bingyang Xiao, and Linjun Wang,* SPADE 1.0: A Simulation Package for Non-Adiabatic Dynamics in Extended Systems, J. Chem. Theory Comput. 21 (7), 3300 (2025). 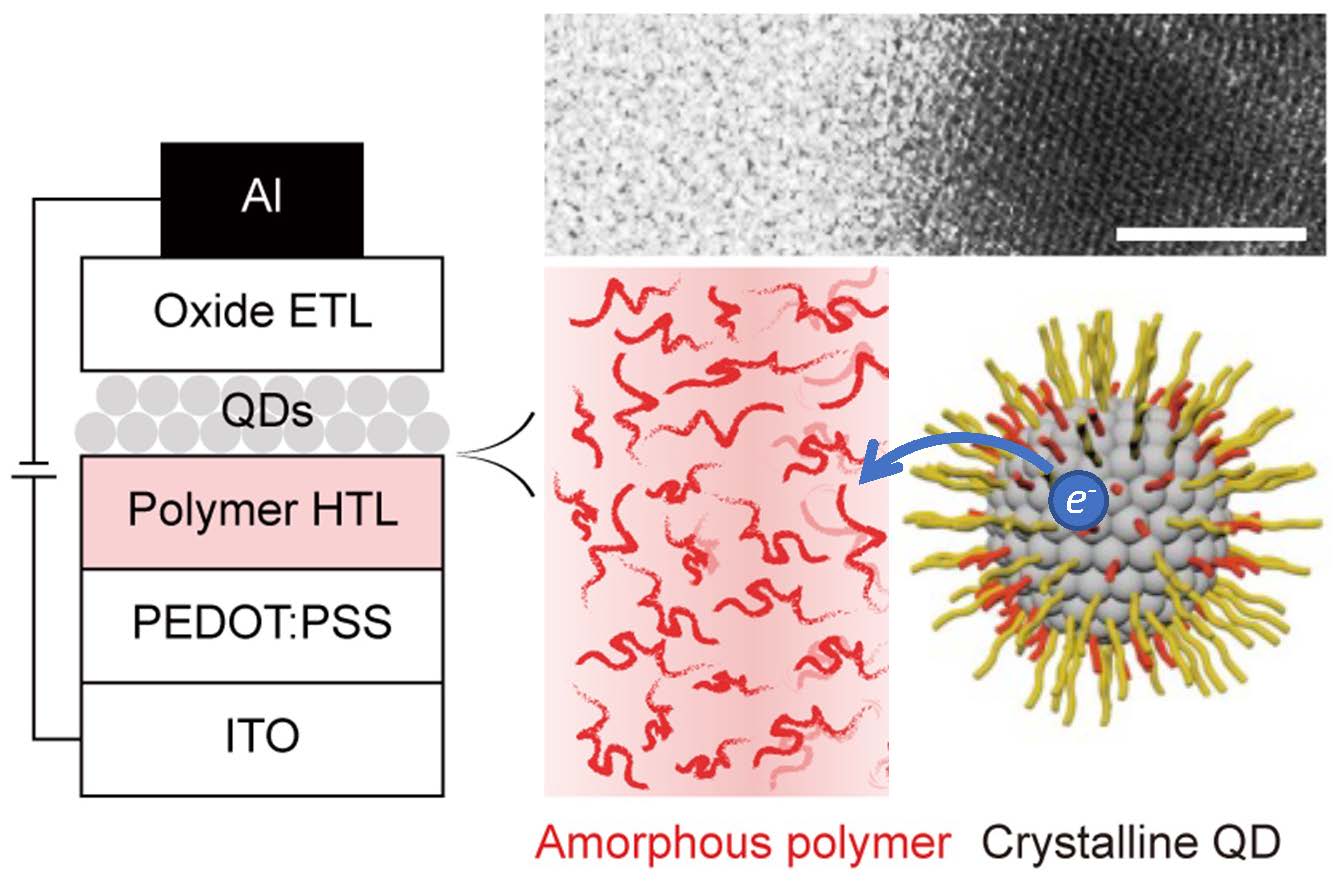 Quantum dots (QDs) are nanoscale semiconductor particles that exhibit unique photoelectronic properties. These properties arise from the phenomenon of quantum confinement, which leads to discrete energy levels within the QDs. We utilize various methods encompassing ab-initio electronic structure calculations, classical molecular dynamics, and mixed quantum-classical nonadiabatic dynamics simulations. Through these approaches, we are able to simulate multi-level processes occurring within QDs, including surface ligand exchange, charge transfer phenomena, as well as their absorption and emission spectra. By employing these comprehensive methodologies, we aim to deepen our understanding of the photoelectronic properties of QDs and pave the way for the development of novel applications. References: 1. Jun Zhang,# Haibing Zhang,# Weicheng Cao,# Zhenfeng Pang, Jiongzhao Li, Yufei Shu, Chenqi Zhu, Xueqian Kong,* Linjun Wang,* and Xiaogang Peng,* Identification of Facet-Dependent Coordination Structures of Carboxylate Ligands on CdSeNanocrystals, J. Am. Chem. Soc. 141 (39), 15675-15683 (2019). 2. Yan Zhang,# Haibing Zhang,# Dongdong Chen, Cheng-Jun Sun, Yang Ren, Jianhui Jiang, Linjun Wang,* Zheng Li,* and Xiaogang Peng,* Engineering of Exciton Spatial Distribution in CdS Nanoplatelets, Nano Lett. 21 (12), 5201-5208 (2021). 3. Jiongzhao Li,# Weicheng Cao,# Yufei Shu,# Haibing Zhang, Xudong Qian, Xueqian Kong,* Linjun Wang,* and Xiaogang Peng,* Water Molecules Bonded to the Carboxylate Groups at the Inorganic-Organic Interface of An Inorganic Nanocrystal Coated with Alkanoate Ligands, Natl. Sci. Rev. 9 (2), nwab138 (2022). 4. Yunzhou Deng,# Feng Peng,# Yao Lu,# Xitong Zhu,# Wangxiao Jin, Jing Qiu, Jiawei Dong, Yanlei Hao, Dawei Di, Yuan Gao, Tulai Sun, Linjun Wang,* Lei Ying,* Fei Huang,* and Yizheng Jin,* Solution-Processed Green and Blue Quantum-Dot Light-Emitting Diodes with Eliminated Charge Leakage, Nat. Photonics 16 (7), 505-511 (2022). 5. Hairui Lei,# Tenghui Li,# Jiongzhao Li, Jie Zhu, Haibing Zhang, Haiyan Qin, Xueqian Kong, Linjun Wang,* and Xiaogang Peng,* Reversible Facet Reconstruction of CdSe/CdS Core/Shell Nanocrystals by Facet-Ligand Pairing, J. Am. Chem. Soc. 145 (12), 6798-6810 (2023). 6. Hairui Lei, Jiongzhao Li,* Xueqian Kong, Linjun Wang,* and Xiaogang Peng,* Toward Surface Chemistry of Semiconductor Nanocrystals at an Atomic-Molecular Level, Acc. Chem. Res. 56 (14), 1966-1977 (2023). 7. Haibing Zhang, Bichuan Cao, Lei Huang, Xiaogang Peng, and Linjun Wang,* Machine Learning Force Field Study of Carboxylate Ligands on the Surface of Zinc-Blende CdSe Quantum Dots, Nano Res. 17 (12), 10685-10693 (2024). 8. Tenghui Li, Jiawei Dong, Zihan Liu, Xiaogang Peng, and Linjun Wang,* Phonon Bottleneck in the Hot Electron Relaxation of n-Doped Quantum Dots: A Large-Scale Nonadiabatic Dynamics Perspective, ACS Nano 20 (7), 5662-5674 (2026).  The goal of global optimization is to find out the structure with the lowest energy. Normally, it is considered as a nondeterministic polynomial-time hard problem because the isomer multiplicity tends to grow exponentially with the number of atoms. We have porposed the fuzzy global optimization (FGO) method. It utilizes mostly the discrete space in a fuzzy search framework. Starting from random initial configurations, we carry out directed Monte Carlo and surface Monte Carlo in the discrete space to obtain low-energy candidate clusters and make real-space local optimizations finally to get the real global minimum structure. FGO is unbiased in nature and has shown high efficiency and reliability in Lennard-Jones, Morse and CdSe clusters. References: 1. Kailiang Yu, Xubo Wang, Liping Chen, and Linjun Wang,* Unbiased Fuzzy Global Optimization of Lennard-Jones Clusters for N ≤ 1000, J. Chem. Phys. 151 (21), 214105 (2019). 2. Liping Chen and Linjun Wang,* Unbiased Fuzzy Global Optimization of Morse Clusters with Short-Range Potential for N ≤ 400, Chin. J. Chem. Phys. 34 (6), 896-904 (2021). 3. Liping Chen, Tao Liang, and Linjun Wang,* Growth Pattern of Large Morse Clusters with Medium-Range Potentials, J. Phys. Chem. Lett. 13 (42), 9801-9808 (2022). 4. Haiwei Lei, Liping Chen,* and Linjun Wang,* Structural Evolution of Cadmium Selenide Clusters: An Unbiased Global Optimization Study of (CdSe)N for 5 ≤ N ≤ 80, J. Phys. Chem. Lett. 14 (25), 5818-5826 (2023). 5. Kaiting Ren,# Tao Liang,# Liping Chen,* and Linjun Wang,* Unbiased Fuzzy Global Optimization of Fullerene Molecular Clusters with the Challenging Girifalco Potential, J. Phys. Chem. Lett. 16 (23) 5855-5861 (2025). 6. Gaoyan Chen,# Tao Liang,# Liping Chen,* and Linjun Wang,* Unbiased Fuzzy Global Optimization with Complex Three-Body Interactions: The Case of Large (C60)N Clusters with the First-Principles PPR Potential, J. Chem. Theory Comput. doi: 10.1021/acs.jctc.6c00329 (2026). |